Last updated: September 26, 2026
Opana ER, Endo Pharmaceuticals’ extended-release oxymorphone product, has no active commercial market in the United States. FDA requested its withdrawal in 2017 after concluding that the product’s abuse by injection created a serious public-health risk. Endo discontinued U.S. sales later that year. The product’s commercial decline had already begun because of opioid-market contraction, payer pressure, generic competition, and litigation exposure. Opana ER now has historical financial value rather than a current revenue opportunity.
What is Opana ER and who marketed it?
Opana ER is the extended-release tablet formulation of oxymorphone hydrochloride, a semi-synthetic opioid analgesic. Endo received FDA approval under NDA 021610 in June 2006 for the management of pain severe enough to require continuous, around-the-clock opioid treatment [1].
| Item |
Detail |
| Brand |
Opana ER |
| Active ingredient |
Oxymorphone hydrochloride |
| Dosage form |
Extended-release oral tablet |
| FDA application |
NDA 021610 |
| Sponsor |
Endo Pharmaceuticals Inc. |
| Initial U.S. approval |
June 2006 |
| Reformulated product |
Introduced in 2012 |
| FDA withdrawal request |
June 2017 |
| U.S. commercial withdrawal |
October 2017 |
| Current U.S. status |
No marketed brand product |
Endo positioned Opana ER against other extended-release opioids, including OxyContin, morphine sulfate ER, fentanyl patches, and hydromorphone ER. Its value proposition was based on oxymorphone’s potency and extended-release delivery rather than a differentiated non-opioid mechanism.
How did Opana ER sales change over time?
Opana ER’s revenue trajectory followed the broader U.S. opioid market: initial growth, peak commercial performance, erosion from policy and payer controls, then an abrupt product exit.
Public filings reported product-level revenue for Opana ER and related oxymorphone products, but reporting conventions changed across periods. Endo’s disclosures also combined or separated Opana ER, Opana immediate-release, and generic oxymorphone products in different filings. That limits direct year-to-year comparisons.
Opana ER commercial timeline
| Period |
Market event |
Financial effect |
| 2006-2011 |
Brand launch and expansion in chronic-pain prescribing |
Revenue growth as an established extended-release opioid |
| 2012 |
Endo launched a reformulated, crush-resistant version |
Temporary lifecycle protection and abuse-deterrence positioning |
| 2013-2014 |
Opioid scrutiny, prescribing restrictions, and payer controls increased |
Slower growth and declining demand |
| 2015-2016 |
Generic and therapeutic substitution pressures intensified |
Lower volume and pricing pressure |
| 2017 |
FDA requested withdrawal because of public-health risks |
End of U.S. brand revenue |
| 2018 onward |
No normal commercial recovery |
Revenue opportunity shifted from sales to litigation, reserves, and restructuring |
Endo’s annual reports described Opana ER as a material product but also identified opioid-related regulatory, litigation, and market risks. The company’s financial exposure extended beyond lost product sales. It included inventory impairment, manufacturing and supply-chain consequences, legal costs, settlement obligations, and the loss of an established controlled-substance franchise [2].
Why did FDA request the withdrawal of Opana ER?
FDA requested removal of Opana ER from the market in June 2017 because the agency concluded that the product’s benefits no longer outweighed its risks when considered in the context of abuse by injection [3].
The decision followed reports that people were manipulating the reformulated tablets for injection. FDA linked intravenous abuse of Opana ER to serious adverse events, including thrombotic thrombocytopenic purpura and kidney injury. The agency also cited an outbreak of serious blood disorders in Tennessee associated with injection of reformulated Opana ER [4].
FDA’s action was unusual because it was based on postmarket public-health evidence involving an abuse-deterrent formulation. The agency did not conclude that reformulation failed to make the tablet more difficult to crush. Instead, it concluded that the reformulated product could still be prepared for injection and that the resulting health consequences were unacceptable.
Endo announced that it would remove the product from the U.S. market. FDA later listed Opana ER as withdrawn for safety or effectiveness reasons, rather than as a product withdrawn solely for commercial reasons [5].
What was the effect of the 2012 reformulation?
Endo introduced a reformulated version of Opana ER intended to resist crushing and manipulation. The reformulation replaced the original formulation and was designed to reduce abuse through snorting or injection.
The reformulation produced two opposing commercial effects:
- It provided a lifecycle-management strategy against generic and competitive pressure.
- It created a new postmarket safety record that ultimately supported FDA’s withdrawal request.
The reformulated product was not an FDA “abuse-deterrent opioid” in the same commercial sense as products that received formal labeling based on abuse-deterrence studies. FDA’s review of Opana ER focused on the actual public-health consequences associated with abuse, particularly injection, rather than on the formulation’s intended physical properties alone [3].
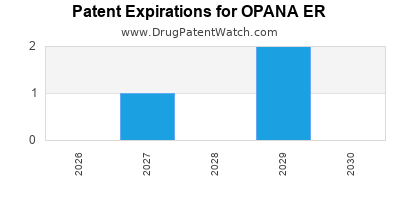
When did Opana ER lose exclusivity and patent protection?
Opana ER’s practical exclusivity ended before the 2017 FDA withdrawal. The product faced regulatory and commercial erosion from generic oxymorphone extended-release products and from substitution into other opioid and non-opioid therapies.
FDA’s Orange Book historically listed patents associated with NDA 021610. Those listings supported patent-certification disputes and generic-launch timing during the product’s commercial life. The precise commercial importance of each patent depended on the dosage strength, formulation, patent certification, litigation outcome, and the timing of FDA approval.
The key point for investors is that patent expiration ceased to be the primary value driver once FDA required removal of the brand product. Even a valid remaining formulation patent would not preserve a U.S. market that the sponsor had exited.
Patent and exclusivity assessment
| Protection category |
Effect on Opana ER |
| FDA approval exclusivity |
Expired before the 2017 withdrawal |
| Original formulation patents |
Provided time-limited protection |
| Reformulation patents |
Supported lifecycle management but did not eliminate abuse risk |
| Method-of-use patents |
Potentially relevant to labeled pain indications, but commercially weak after withdrawal |
| Orange Book listings |
Relevant to generic certifications before market exit |
| Current market value |
Minimal because there is no ongoing U.S. brand sales platform |
No current U.S. product strategy can rely on Opana ER’s historical patent estate alone. The product’s principal barrier became regulatory and reputational risk, not patent scarcity.
Were there Paragraph IV challenges to Opana ER?
Generic applicants for branded drug products can file Paragraph IV certifications alleging that listed patents are invalid, unenforceable, or not infringed. Opana ER was subject to the type of generic competition and patent-certification activity expected for a high-value extended-release opioid.
The commercial consequences were material:
- Generic entry reduced brand pricing power.
- Multiple strengths increased substitution opportunities.
- Extended-release opioids faced declining prescription volume even before FDA withdrawal.
- Litigation and settlement terms could delay or accelerate generic entry.
- A generic product could remain commercially viable after the brand’s withdrawal, subject to FDA approval and market demand.
Publicly available company and FDA materials do not support treating Paragraph IV litigation as the principal reason for Opana ER’s ultimate market exit. FDA’s safety determination, not a patent judgment, ended the brand’s U.S. commercial program.
What is the Orange Book status of Opana ER?
Opana ER is not an active marketed brand opportunity in the U.S. Orange Book. FDA’s withdrawal action was based on safety and effectiveness concerns, and the product was removed from the market after Endo discontinued it [5].
The distinction matters:
- A drug may be listed historically in Orange Book records.
- A listed patent may remain relevant to past generic litigation.
- A product can lose commercial value even before every associated patent expires.
- FDA withdrawal for safety reasons is materially different from ordinary discontinuation for low sales.
Generic oxymorphone products, where approved, do not recreate the original Opana ER franchise. They operate within a heavily restricted opioid market and face demand, prescribing, reimbursement, and regulatory barriers.
Which companies challenged or competed with Opana ER?
Opana ER competed with branded and generic products across the extended-release opioid segment.
Principal competitive products
| Competitor |
Active ingredient |
Competitive position |
| OxyContin |
Oxycodone hydrochloride ER |
Large branded extended-release opioid franchise |
| Xtampza ER |
Oxycodone ER |
Abuse-deterrent formulation with later market entry |
| Hysingla ER |
Hydrocodone ER |
Abuse-deterrent extended-release opioid |
| Exalgo |
Hydromorphone ER |
High-potency extended-release opioid |
| Morphine sulfate ER |
Morphine |
Lower-cost generic alternative |
| Fentanyl transdermal systems |
Fentanyl |
Non-oral extended-release delivery option |
| Generic oxymorphone ER |
Oxymorphone hydrochloride ER |
Direct generic substitute |
Competition was not limited to branded opioids. Formularies increasingly favored lower-cost generics, while clinical guidelines and state policies reduced use of long-term opioid therapy. This weakened the entire category.
What litigation affected Opana ER and Endo?
Opana ER was part of the broader opioid litigation environment involving manufacturers, distributors, prescribers, and pharmacies. Claims generally alleged that opioid companies overstated benefits, minimized addiction risks, failed to control suspicious orders, or contributed to widespread opioid misuse.
Endo disclosed substantial opioid-related litigation and government investigations in its SEC filings. Opana ER was also connected to product-liability and marketing-related claims. The FDA withdrawal increased litigation exposure because plaintiffs could use the agency’s public-health findings to support allegations concerning abuse, risk management, and product design [2].
The financial effect was broader than a single settlement payment. Endo faced:
- Legal defense costs.
- Settlement contributions.
- Insurance and indemnity disputes.
- Potential damages and governmental claims.
- Reduced value of opioid-related assets.
- Restructuring pressure across the company.
Endo filed for Chapter 11 bankruptcy protection in August 2022, citing debt burdens, opioid litigation, and operating pressures. The bankruptcy process materially reduced the likelihood that Opana ER could generate recoverable value for unsecured stakeholders independent of broader restructuring outcomes [6].
How strong was the Opana ER patent estate?
The patent estate was commercially effective during the product’s growth phase but weak as a long-term defense after withdrawal.
Strengths
- Extended-release formulation patents supported a branded lifecycle.
- Reformulation delayed simple substitution and provided additional product differentiation.
- Orange Book listings created potential litigation leverage against generic applicants.
- Oxymorphone’s potency and dosage-form characteristics supported physician familiarity.
Weaknesses
- Patent protection could not address injection abuse.
- FDA safety action outweighed remaining exclusivity value.
- Opioid prescribing restrictions reduced the addressable market.
- Generic and therapeutic substitutes limited pricing power.
- Product withdrawal eliminated the value of future branded sales.
For valuation purposes, Opana ER should be classified as a discontinued commercial asset with historical patent relevance, not as an active protected franchise.
What generic launch risks existed?
Generic launch risk was high before the brand withdrawal and changed form afterward.
Before withdrawal
A generic entrant could cause rapid price erosion because:
- Payers commonly prefer generic extended-release opioids.
- The active ingredient was chemically established.
- Physicians and pharmacies had existing substitution pathways.
- Opana ER faced competition from multiple opioid classes.
- The market was already declining because of prescribing controls.
After withdrawal
Generic manufacturers faced a different risk profile:
- FDA safety concerns remained associated with oxymorphone ER.
- Demand for extended-release opioids was structurally lower.
- State-level opioid restrictions constrained volume.
- Product-liability exposure could affect supply decisions.
- Reimbursement might not justify manufacturing and compliance costs.
The likely generic-launch scenario was therefore limited-volume substitution, not restoration of the original brand market.
What was the financial trajectory for Endo?
Opana ER contributed to Endo’s historical product portfolio but could not offset the company’s later debt, litigation, and operating problems. The product’s financial trajectory had four stages.
Growth and portfolio contribution
Following FDA approval, Opana ER expanded Endo’s branded pain portfolio and created recurring prescription revenue. The product benefited from the broader growth of extended-release opioid prescribing.
Erosion
Sales weakened as regulators, payers, and prescribers reassessed chronic opioid therapy. Generic competition and lower opioid utilization reduced both volume and pricing.
Forced withdrawal
The 2017 FDA action ended the product’s U.S. revenue stream. The reformulation, which had been intended to extend the lifecycle, became part of the agency’s risk analysis.
Litigation and restructuring
After withdrawal, Opana ER’s economic significance shifted from operating revenue to liability exposure. Endo’s opioid obligations formed part of the company’s broader financial distress and Chapter 11 restructuring [6].
The product should not be modeled as a future growth asset. Any valuation should treat historical Opana ER revenue as nonrecurring and assign no base-case U.S. brand sales after 2017.
How does Opana ER compare with OxyContin and Xtampza ER?
| Factor |
Opana ER |
OxyContin |
Xtampza ER |
| Active ingredient |
Oxymorphone |
Oxycodone |
Oxycodone |
| U.S. status |
Withdrawn |
Marketed, subject to restrictions |
Marketed |
| Abuse-deterrent strategy |
Reformulated tablet |
Reformulated tablet |
Abuse-deterrent formulation |
| FDA safety outcome |
Withdrawal requested in 2017 |
Continued marketing under restrictions |
Continued marketing |
| Current commercial value |
Historical |
Ongoing but constrained |
Ongoing but constrained |
| Generic risk |
High historically |
High, with complex formulation issues |
Future generic risk remains |
| Litigation exposure |
Significant |
Significant |
Opioid-sector exposure |
The comparison shows why abuse-deterrent formulation alone did not guarantee commercial durability. FDA evaluated Opana ER against observed public-health outcomes, not only laboratory manipulation resistance.
What is the current investment and licensing outlook for Opana ER?
The licensing case is weak. A buyer would acquire historical know-how, trademarks, regulatory records, and possibly intellectual property, but not a normal branded opioid revenue stream.
Potential asset value would be limited to:
- Non-U.S. rights, if any remain active.
- Historical formulation and manufacturing know-how.
- Patent rights with remaining term in selected jurisdictions.
- Data and regulatory documentation.
- Litigation or settlement-related claims, subject to court treatment.
A relaunch in the United States would face major regulatory obstacles. It would require a new commercial strategy, extensive abuse-risk controls, manufacturing compliance, and a persuasive public-health case against the record that led to withdrawal. The economic return would be difficult to justify against generic competition and opioid-market contraction.
Key Takeaways
- Opana ER was approved in 2006 and withdrawn from the U.S. market in 2017 after FDA concluded that its abuse by injection created serious public-health risks.
- Endo’s 2012 reformulation delayed ordinary product erosion but did not prevent FDA withdrawal.
- Patent and Orange Book protection became commercially secondary once the product was removed for safety reasons.
- Generic and therapeutic competition, opioid prescribing restrictions, and payer controls had already weakened the franchise.
- Opana ER has no meaningful current U.S. brand revenue base.
- Endo’s opioid litigation and financial distress, including its 2022 Chapter 11 filing, reduced the economic value of the product’s legacy assets.
- The product is best valued as a discontinued pharmaceutical asset with historical patent and litigation significance.
FAQs
Is Opana ER still available in the United States?
No. Endo discontinued U.S. sales after FDA requested withdrawal in 2017.
Can a generic company still sell oxymorphone ER?
FDA approval and commercial availability are separate issues. A generic applicant may obtain approval subject to FDA requirements, but market demand, safety concerns, opioid controls, and product-liability exposure materially restrict commercial potential.
Did FDA withdraw Opana ER because the patent expired?
No. FDA’s action was based on safety and public-health concerns associated with abuse by injection.
Does Opana ER have biosimilar competition?
No. Opana ER is a chemically synthesized small-molecule drug, not a biologic. The relevant competitors are generic oxymorphone products and other opioid or non-opioid analgesics.
Could Opana ER be relaunched after Endo’s bankruptcy?
A relaunch would require regulatory, manufacturing, commercial, and litigation solutions that do not exist through patent ownership alone. The historical U.S. franchise should not be assumed to be recoverable.
References
-
U.S. Food and Drug Administration. (2006). Opana ER prescribing information and approval history. FDA.
-
Endo International plc. (2016). Annual report on Form 10-K for the fiscal year ended December 31, 2016. U.S. Securities and Exchange Commission.
-
U.S. Food and Drug Administration. (2017, June 8). FDA requests removal of Opana ER for risks related to abuse. FDA.
-
U.S. Food and Drug Administration. (2017). FDA briefing document: Joint meeting of the Anesthetic and Analgesic Drug Products Advisory Committee and the Drug Safety and Risk Management Advisory Committee. FDA.
-
U.S. Food and Drug Administration. (2018). Approved drug products with therapeutic equivalence evaluations: Orange Book. FDA.
-
Endo International plc. (2022). Current report on Form 8-K announcing voluntary Chapter 11 proceedings. U.S. Securities and Exchange Commission.